
Adenylosuccinate lyase (ADSL) deficiency (OMIM 103050; ORPHA:46) is a rare autosomal recessive disorder of purine metabolism caused by biallelic pathogenic variants in the ADSL gene. It primarily affects the central nervous system and shows a broad clinical spectrum ranging from fatal neonatal encephalopathy to mild neurodevelopmental phenotypes.
Clinical spectrum
ADSL deficiency presents as a continuous clinical spectrum, historically described as a prenatal form, a fatal neonatal form, a severe form (type I), and a mild to moderate form (type II). The main manifestations are developmental delay or intellectual disability, epilepsy, hypotonia, impaired speech and communication, autistic or behavioral features, microcephaly, movement abnormalities including ataxia, tremor or dystonia, and visual abnormalities such as strabismus, nystagmus and impaired visual contact. Severe neonatal cases may additionally present with fetal hypokinesia, feeding difficulties, apnea and respiratory failure. At the mild end of the spectrum, some patients may have only mild developmental or intellectual impairment without epilepsy or structural brain abnormalities. Considerable phenotypic variability may occur, including among patients within the same family.
Biochemical basis
ADSL (EC 4.3.2.2) is a bifunctional enzyme involved in both de novo purine synthesis (DNPS) and the purine nucleotide cycle. It catalyzes the conversion of succinylaminoimidazole carboxamide ribotide (SAICAR) to aminoimidazole carboxamide ribotide (AICAR) and fumarate, and the conversion of adenylosuccinate (SAMP) to AMP and fumarate.
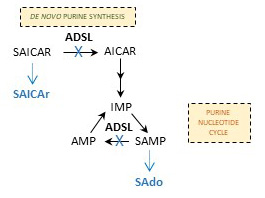 ADSL deficiency leads to accumulation of the dephosphorylated succinylpurines succinylaminoimidazole carboxamide riboside (SAICAr) and succinyladenosine (SAdo). These metabolites can be detected in urine, plasma and cerebrospinal fluid and represent characteristic biochemical markers of the disease.
The mechanisms responsible for the neurological phenotype and the marked clinical heterogeneity are not yet fully understood. In addition to succinylpurine accumulation, ADSL deficiency affects purine homeostasis, metabolic flux through de novo purine synthesis and purinosome assembly. Absolute SAdo and SAICAr concentrations do not reliably predict clinical severity. An association between the SAdo/SAICAr ratio in cerebrospinal fluid and disease severity has been reported, but this ratio should not be considered a general prognostic marker, particularly in urine.
Diagnosis
The diagnostic pathway has changed substantially with the widespread use of next-generation sequencing. Today, many patients are initially identified by whole-exome sequencing (WES), genome sequencing or multigene panels, because the clinical phenotype of ADSL deficiency is nonspecific. Identification of biallelic pathogenic or likely pathogenic variants in the ADSL gene should, whenever feasible, be complemented by biochemical confirmation through measurement of SAdo and SAICAr in urine and/or plasma.
Conversely, detection of elevated SAdo and SAICAr may lead directly to targeted molecular analysis of the ADSL gene. Functional studies, including measurement of ADSL enzyme activity, may provide additional evidence for interpretation of variants of uncertain significance.
ADSL variants and patients
Our ADSL patient database collects information on published patients with ADSL deficiency. A separate ADSL variant database provides an overview of reported disease-associated ADSL variants.
Treatment
Clinical management of ADSL deficiency remains primarily symptomatic, including treatment of epilepsy and multidisciplinary neurodevelopmental care. Disease-modifying therapeutic approaches are currently under investigation.
SAdo and SAICAr standards
We provide SAdo and SAICAr standards, including isotopically labelled compounds, for diagnostic and research laboratories.
Literature
For current publications on adenylosuccinate lyase deficiency, see PubMed – adenylosuccinate lyase deficiency.